



[ad_1]
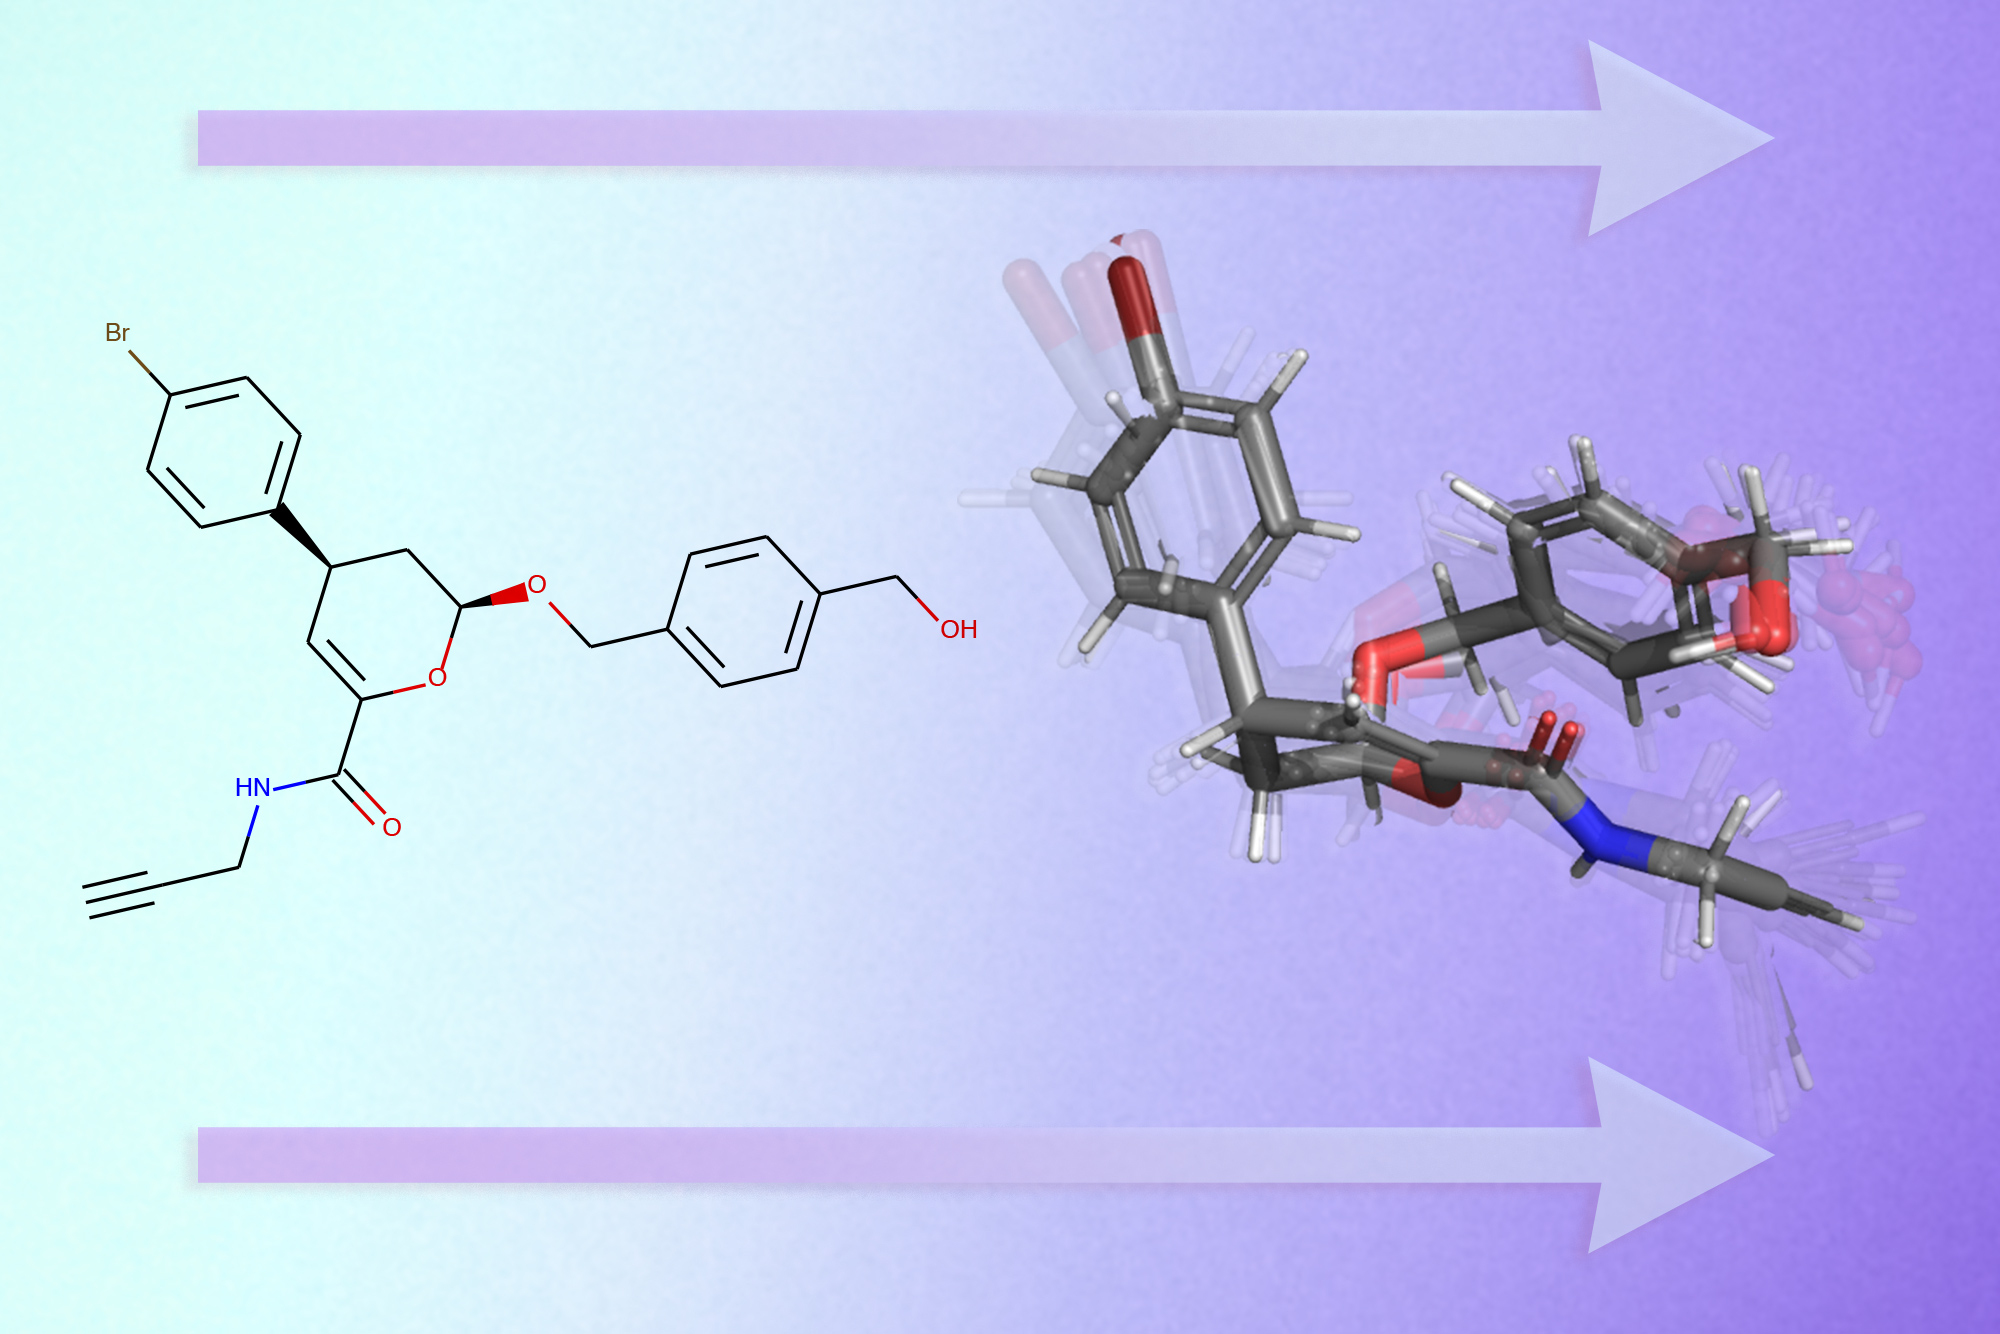
Of their quest to find efficient new medicines, scientists seek for drug-like molecules that may connect to disease-causing proteins and alter their performance. It’s essential that they know the 3D form of a molecule to know the way it will connect to particular surfaces of the protein.
However a single molecule can fold in hundreds of various methods, so fixing that puzzle experimentally is a time consuming and costly course of akin to looking for a needle in a molecular haystack.
MIT researchers are utilizing machine studying to streamline this advanced process. They’ve created a deep studying mannequin that predicts the 3D shapes of a molecule solely based mostly on a graph in 2D of its molecular construction. Molecules are sometimes represented as small graphs.
Their system, GeoMol, processes molecules in solely seconds and performs higher than different machine studying fashions, together with some industrial strategies. GeoMol may assist pharmaceutical corporations speed up the drug discovery course of by narrowing down the variety of molecules they should check in lab experiments, says Octavian-Eugen Ganea, a postdoc within the Laptop Science and Synthetic Intelligence Laboratory (CSAIL) and co-lead creator of the paper.
“When you’re fascinated by how these constructions transfer in 3D area, there are actually solely sure components of the molecule which might be truly versatile, these rotatable bonds. One of many key improvements of our work is that we take into consideration modeling the conformational flexibility like a chemical engineer would. It’s actually about attempting to foretell the potential distribution of rotatable bonds within the construction,” says Lagnajit Pattanaik, a graduate pupil within the Division of Chemical Engineering and co-lead creator of the paper.
Different authors embody Connor W. Coley, the Henri Slezynger Profession Growth Assistant Professor of Chemical Engineering; Regina Barzilay, the College of Engineering Distinguished Professor for AI and Well being in CSAIL; Klavs F. Jensen, the Warren Okay. Lewis Professor of Chemical Engineering; William H. Inexperienced, the Hoyt C. Hottel Professor in Chemical Engineering; and senior creator Tommi S. Jaakkola, the Thomas Siebel Professor of Electrical Engineering in CSAIL and a member of the Institute for Information, Techniques, and Society. The analysis shall be introduced this week on the Convention on Neural Info Processing Techniques.
Mapping a molecule
In a molecular graph, a molecule’s particular person atoms are represented as nodes and the chemical bonds that join them are edges.
GeoMol leverages a latest instrument in deep studying referred to as a message passing neural community, which is particularly designed to function on graphs. The researchers tailored a message passing neural community to foretell particular components of molecular geometry.
Given a molecular graph, GeoMol initially predicts the lengths of the chemical bonds between atoms and the angles of these particular person bonds. The way in which the atoms are organized and linked determines which bonds can rotate.
GeoMol then predicts the construction of every atom’s native neighborhood individually and assembles neighboring pairs of rotatable bonds by computing the torsion angles after which aligning them. A torsion angle determines the movement of three segments which might be linked, on this case, three chemical bonds that join 4 atoms.
“Right here, the rotatable bonds can take an enormous vary of potential values. So, the usage of these message passing neural networks permits us to seize quite a lot of the native and international environments that influences that prediction. The rotatable bond can take a number of values, and we wish our prediction to have the ability to mirror that underlying distribution,” Pattanaik says.
Overcoming current hurdles
One main problem to predicting the 3D construction of molecules is to mannequin chirality. A chiral molecule can’t be superimposed on its mirror picture, like a pair of arms (regardless of the way you rotate your arms, there isn’t any means their options precisely line up). If a molecule is chiral, its mirror picture received’t work together with the atmosphere in the identical means.
This might trigger medicines to work together with proteins incorrectly, which may end in harmful uncomfortable side effects. Present machine studying strategies typically contain an extended, advanced optimization course of to make sure chirality is appropriately recognized, Ganea says.
As a result of GeoMol determines the 3D construction of every bond individually, it explicitly defines chirality in the course of the prediction course of, eliminating the necessity for optimization after-the-fact.
After performing these predictions, GeoMol outputs a set of possible 3D constructions for the molecule.
“What we are able to do now’s take our mannequin and join it end-to-end with a mannequin that predicts this attachment to particular protein surfaces. Our mannequin is just not a separate pipeline. It is rather straightforward to combine with different deep studying fashions,” Ganea says.
A “super-fast” mannequin
The researchers examined their mannequin utilizing a dataset of molecules and the possible 3D shapes they might take, which was developed by Rafael Gomez-Bombarelli, the Jeffrey Cheah Profession Growth Chair in Engineering, and graduate pupil Simon Axelrod.
They evaluated what number of of those possible 3D constructions their mannequin was in a position to seize, compared to machine studying fashions and different strategies.
In practically all cases, GeoMol outperformed the opposite fashions on all examined metrics.
“We discovered that our mannequin is super-fast, which was actually thrilling to see. And importantly, as you add extra rotatable bonds, you count on these algorithms to decelerate considerably. However we didn’t actually see that. The velocity scales properly with the variety of rotatable bonds, which is promising for utilizing all these fashions down the road, particularly for purposes the place you are attempting to shortly predict the 3D constructions inside these proteins,” Pattanaik says.
Sooner or later, the researchers hope to use GeoMol to the realm of high-throughput digital screening, utilizing the mannequin to find out small molecule constructions that will work together with a selected protein. Additionally they wish to hold refining GeoMol with extra coaching information so it may possibly extra successfully predict the construction of lengthy molecules with many versatile bonds.
“Conformational evaluation is a key part of quite a few duties in computer-aided drug design, and an essential part in advancing machine studying approaches in drug discovery,” says Pat Walters, senior vice chairman of computation at Relay Therapeutics, who was not concerned on this analysis. “I’m excited by persevering with advances within the area and thank MIT for contributing to broader learnings on this space.”
This analysis was funded by the Machine Studying for Pharmaceutical Discovery and Synthesis consortium.
[ad_2]

